Test Categories: IVD, LDT, ASR, RUO
Where a molecular test comes from decides what the lab must prove: FDA-cleared or approved kits, modified kits and lab-developed tests, analyte-specific reagents, and research- and investigational-use products - plus where FDA's LDT rule stands.
- 4 min
- 7 steps
- 3 questions
- Lesson 41 of 60
In this lesson
- Why the category matters
- FDA-cleared and approved tests
- Laboratory-developed tests
- Analyte-specific reagents
- RUO and IUO
- CLIA complexity
- What to take from this
Picking up where you left off.
Why the category matters
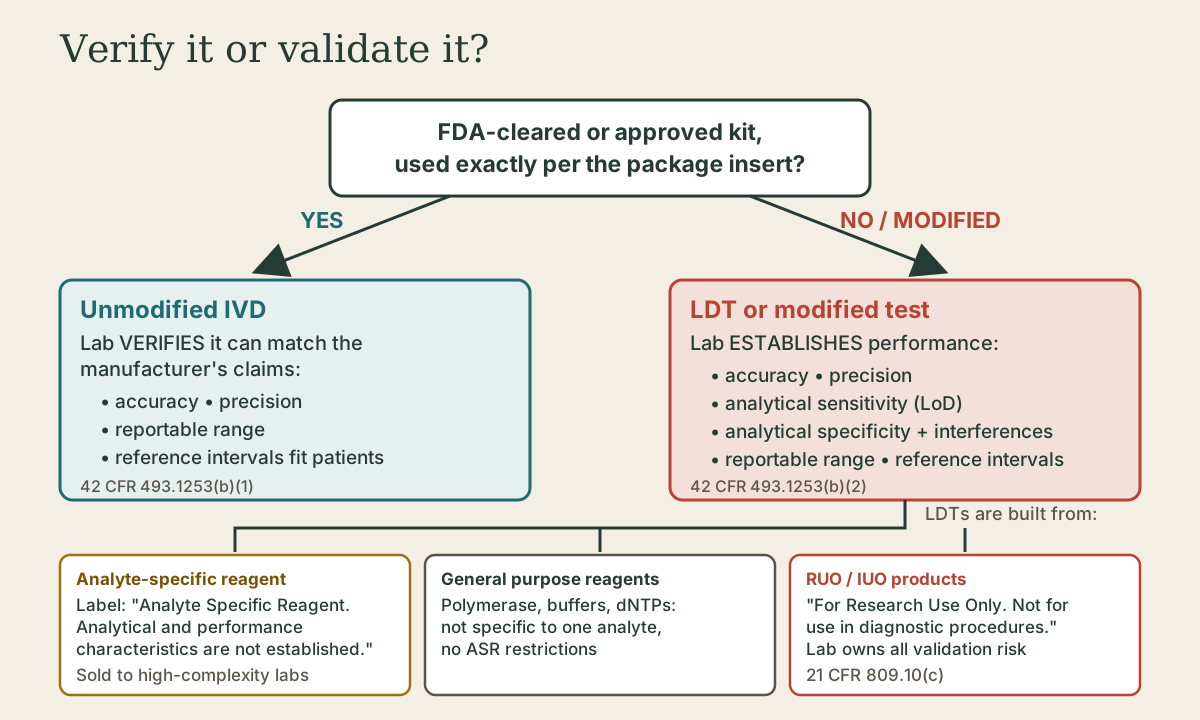
What a lab must do before reporting patient results depends on where the test came from. Under CLIA, an unmodified FDA-cleared or approved test needs verification; a modified test or a laboratory-developed test needs full validation 1. The next lesson covers both. First, the categories.
FDA-cleared and approved tests
An in vitro diagnostic (IVD) kit that FDA has cleared (510(k) or De Novo) or approved (premarket approval, for higher-risk tests such as many companion diagnostics) comes with a package insert defining its intended use, specimen types, procedure, and performance.
Used exactly as labeled, it’s an unmodified IVD. The lab verifies it can reproduce the manufacturer’s accuracy, precision, reportable range, and reference intervals 1.
Changing anything material makes it a modified FDA test:
- A different specimen type (rectal swabs on an assay cleared for urine).
- A different extraction method or instrument.
- Changed volumes, cycling, or cutoffs.
- A different patient population or intended use.
Modified tests are treated like LDTs: the lab must establish performance 1.
Quick check
Changing the specimen type, procedure, or intended use modifies the cleared test. The lab must establish performance specifications as it would for an LDT.
Laboratory-developed tests
A laboratory-developed test (LDT) is designed, validated, and used within a single CLIA-certified high-complexity lab. Much molecular testing is LDTs: rare-disease sequencing, many oncology panels, MRD assays.
For decades FDA claimed authority over LDTs but used enforcement discretion. In May 2024 it issued a rule to phase in device requirements for LDTs. On March 31, 2025, the U.S. District Court for the Eastern District of Texas vacated that rule in ACLA v. FDA and AMP v. FDA, holding that LDTs aren’t “devices” under the Food, Drug, and Cosmetic Act 2. FDA formally rescinded the rule effective September 19, 2025 3. LDTs remain regulated through CLIA (and states such as New York).
Analyte-specific reagents
An analyte-specific reagent (ASR) is a building block - an antibody, a specific primer or probe set - sold for labs to build their own tests. Rules 4:
- ASRs may be sold only to IVD manufacturers, CLIA high-complexity laboratories, and organizations using them for non-diagnostic purposes.
- Labels read: “Analyte Specific Reagent. Analytical and performance characteristics are not established.”
- A test built in-house with ASRs carries a report statement that the test “was developed and its performance characteristics determined by (Laboratory Name). It has not been cleared or approved by the U.S. Food and Drug Administration.”
General purpose reagents (buffers, polymerases, nucleotides) aren’t specific to an analyte and carry no such restrictions.
Quick check
21 CFR 809.30 sets the ASR label statement. ASRs may be sold only to IVD manufacturers, CLIA high-complexity labs, and non-diagnostic users.
RUO and IUO
- Research Use Only (RUO): labeled “For Research Use Only. Not for use in diagnostic procedures.” 5. Instruments and reagents sold for research aren’t meant for clinical testing. A lab that does use an RUO component in a clinical LDT takes on full responsibility for validating it, and regulators and accreditors look hard at the practice.
- Investigational Use Only (IUO): products in clinical investigation, labeled “For Investigational Use Only. The performance characteristics of this product have not been established.” 5.
CLIA complexity
Separately, FDA categorizes tests by complexity for CLIA: waived, moderate, or high. Most molecular tests are high complexity, and all LDTs are high complexity by default. A growing number of simple sample-to-answer molecular tests are CLIA-waived (the first molecular waiver came in 2015), which is moving molecular testing into clinics and pharmacies.
What to take from this
An unmodified FDA-cleared or approved test is verified; a modified test or LDT must be fully validated. ASRs are sold only to manufacturers, high-complexity labs, and non-diagnostic users, with a fixed label and a report disclaimer for in-house tests. RUO and IUO products carry their own label statements and aren’t intended for routine diagnosis. FDA’s 2024 LDT rule was vacated in March 2025 and rescinded that September, so LDTs remain under CLIA.
Practice
The Eastern District of Texas vacated the rule on March 31, 2025, holding LDTs aren’t devices under the FD&C Act; FDA rescinded it effective September 19, 2025. LDTs remain under CLIA.
Lesson complete
Nice work.
Sources for this lesson
- 142 CFR 493.1253 - Establishment and verification of performance specifications. Legal Information Institute (CLIA regulations). verifiedUnmodified FDA-cleared systems - verify accuracy, precision, reportable range, reference intervals; modified systems and LDTs - establish those plus analytical sensitivity and analytical specificity including interfering substances. Cited at: (b)(1); (b)(2).
- 2Federal District Court Vacates FDA's Laboratory Developed Tests Final Rule. FDA Law Blog (Hyman, Phelps & McNamara). 2025. verifiedOn March 31, 2025, the E.D. Texas vacated FDA's 2024 rule treating LDTs as devices (ACLA v. FDA; AMP v. FDA); FDA formally rescinded the rule effective September 19, 2025.
- 3FDA Withdraws Final Rule Regulating Lab-Developed Tests as Medical Devices. Clinical Lab Products. 2025. verifiedFDA rescinded the 2024 LDT rule in September 2025, returning the IVD regulation to its earlier wording.
- 421 CFR 809.30 - Restrictions on the sale, distribution and use of analyte specific reagents. Legal Information Institute. verifiedASRs sold to IVD manufacturers, CLIA high-complexity labs, and non-diagnostic users; labeled "Analyte Specific Reagent. Analytical and performance characteristics are not established."; in-house test reports carry the "developed and its performance characteristics determined by" disclaimer.
- 521 CFR 809.10 - Labeling for in vitro diagnostic products. Legal Information Institute. verifiedRUO - 'For Research Use Only. Not for use in diagnostic procedures.' IUO - 'For Investigational Use Only. The performance characteristics of this product have not been established.' Cited at: (c)(2)(i); (c)(2)(ii).